resources
Integrated Medical Device Manufacturing for OEMs: From Engineering to Supply Chain

Published on 26th Sept 2026
Medical device OEMs are under pressure from every direction. Regulatory requirements are tightening. Time-to-market windows are shrinking. And the cost of a quality failure, whether it is a non-conforming component or a documentation gap that triggers an FDA audit, is measured not just in dollars but in patient safety and brand reputation.
The traditional response has been to spread manufacturing across multiple specialist vendors: one for injection moulding, another for electronics, a third for assembly, and a fourth for testing. On paper, this looks like risk diversification. In practice, it multiplies handoffs, fragments accountability, and introduces quality gaps at every transition point.
The global medical device contract manufacturing market reached USD 105.53 billion in 2026 and is projected to grow to USD 171 billion by 2031, according to Mordor Intelligence. The fastest-growing segment is device development and manufacturing services, which already commands 53.72% of market share. The reason is straightforward: OEMs are consolidating their supply chains around integrated partners who can absorb complexity rather than distribute it.
This article covers what a truly integrated medical device manufacturing partnership looks like, what capabilities matter at each stage of the OEM journey, and how FVG delivers that model across engineering, precision moulding, electronics, assembly, testing, and supply-chain management.
Most procurement teams inherit a multi-vendor model because it evolved organically: the best moulding shop in one region, the most competitive PCB assembler in another, a testing house that was added when the original partner lacked the capability. The result is a supply chain that is optimised for individual cost lines but not for overall programme performance.
The hidden costs are significant.
Every time a component or sub-assembly moves between vendors, three things happen: traceability documentation must be transferred, quality responsibility shifts, and lead time accumulates. In a regulated environment, each handoff is also a potential audit finding. FDA 21 CFR Part 820 requires that OEMs maintain documented control over their entire supply chain, including all sub-tier suppliers. When your moulding vendor uses a sub-contracted tooling house, that sub-contractor is part of your quality obligation, whether you know it or not.
Multi-vendor programmes create a predictable dynamic: when something goes wrong, each vendor points to the previous one. The moulding supplier says the material specification was ambiguous. The electronics assembler says the substrate was out of tolerance. The testing house says the failure mode was introduced upstream. Meanwhile, the OEM's programme manager is mediating disputes instead of managing the programme.
A study of medical device outsourcing practices found that capacity misalignment between vendors is one of the most common causes of programme delays. When your moulding partner's lead time extends by three weeks, every downstream vendor's schedule shifts. With a single integrated partner, that schedule risk is internal and managed proactively.
The real risk of vendor fragmentation is not cost. It is the regulatory and timeline exposure that accumulates invisibly across every handoff.
The term "integrated" is used loosely in contract manufacturing. Many providers claim integration while still relying heavily on sub-contractors for core process steps. True integration means that every critical manufacturing process, from material selection through final test, is performed under a single quality management system, within a single accountability structure, and with a single point of contact for the OEM.
For medical device programmes, this matters for one overriding reason: your quality system is only as strong as your weakest supplier's quality system. When your contract manufacturer is genuinely integrated, their ISO 13485 certification covers the entire scope of work, not just the steps they perform directly.
A genuinely integrated medical device manufacturing partner covers all of the following:
Capability Area | What to Verify |
|---|---|
Engineering & DFM | In-house engineering team, design-for-manufacturability review, prototyping capability |
Materials & Tooling | Medical-grade material selection, in-house tooling design and maintenance |
Precision Moulding | Cleanroom injection moulding, validated processes, IQ/OQ/PQ documentation |
Electronics (PCBA) | SMT assembly, IPC-A-610 compliance, traceability to component level |
Box Build & Assembly | Electromechanical integration, ESD-controlled environments |
Testing & Inspection | Functional test, AOI, X-ray, IEC 60601 compliance testing |
Supply Chain Management | Approved vendor lists, incoming inspection, inventory management |
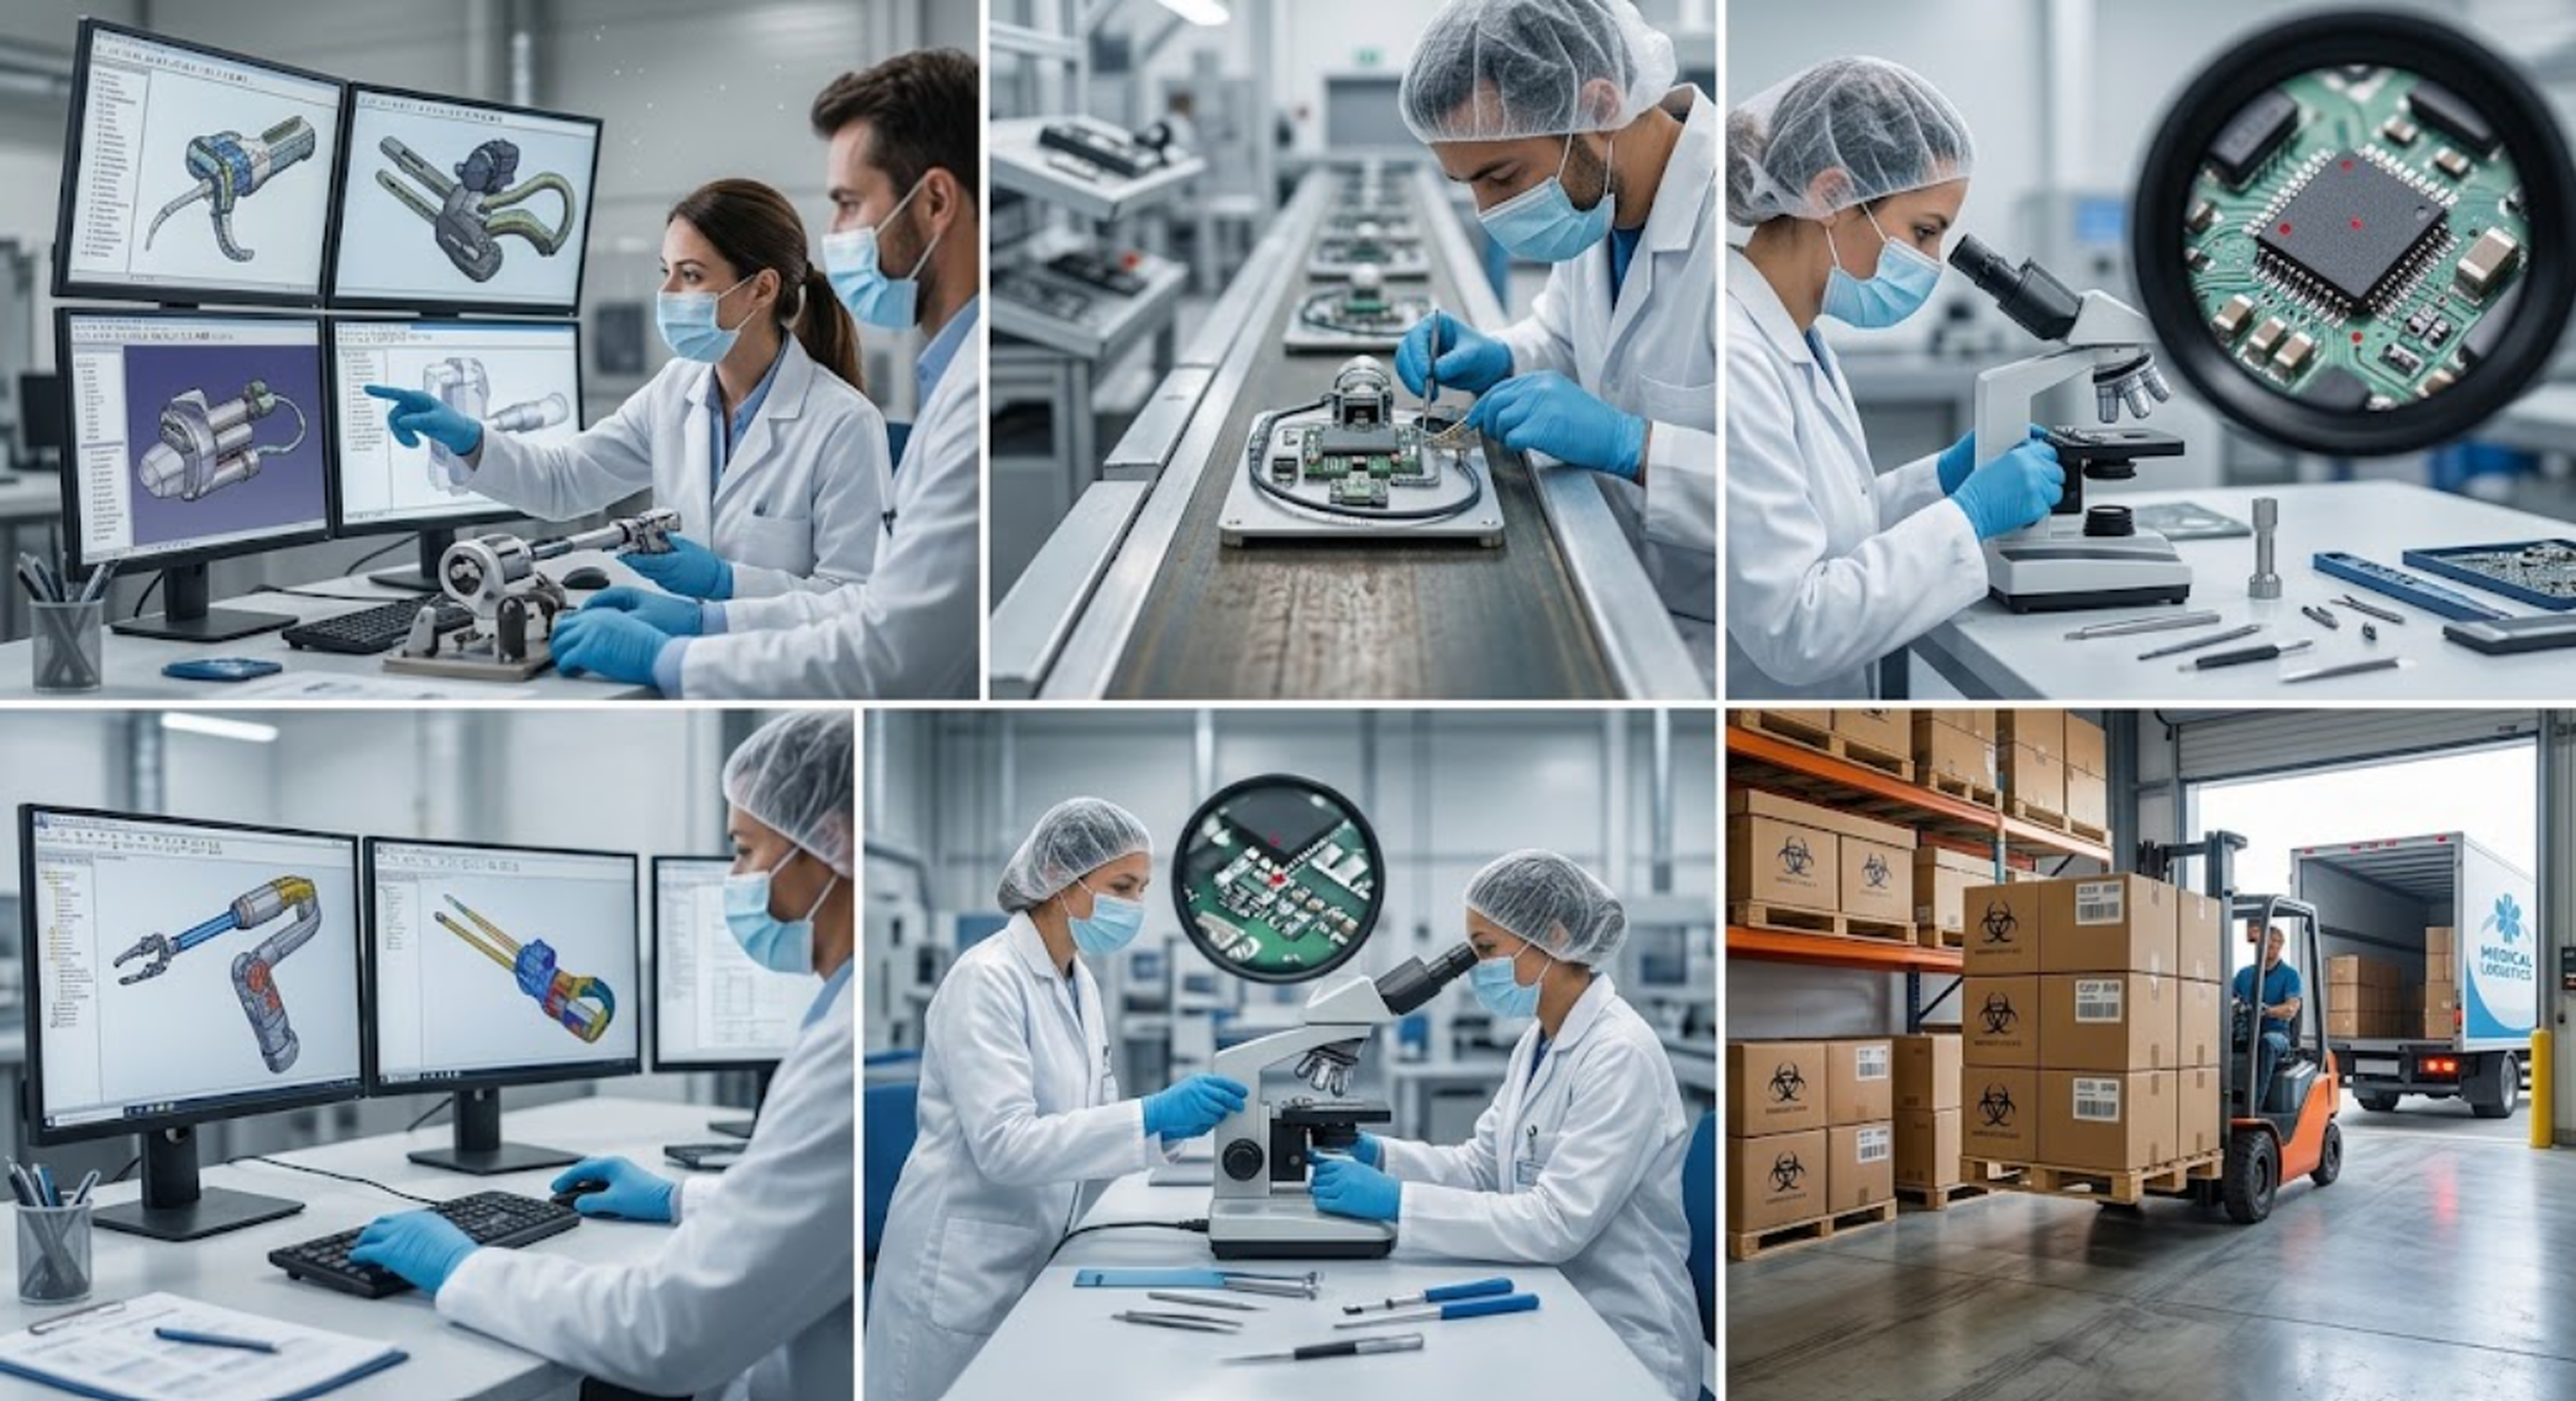
FVG operates across every layer of this stack. With over 30 years of engineering expertise and facilities spanning Asia and North America, FVG functions as a vertically integrated Electronics Manufacturing Services provider for medical device OEMs at low-to-medium volume, high-mix production profiles.
Most manufacturing failures in medical devices are not manufacturing failures at all. They are design failures that only become visible during production. A component geometry that is theoretically correct but practically unmoulable. A material choice that passes bench testing but degrades under sterilisation conditions. A PCB layout that meets electrical specifications but creates assembly defects at volume.
This is why engineering engagement at the front end of a programme is not a nice-to-have. It is the single most effective risk mitigation available to an OEM procurement team.
DFM for medical devices goes beyond the standard considerations of draft angles, wall thickness, and gate placement. It must account for:
Cleanroom compatibility: Can the part geometry be moulded and assembled in an ISO Class 7 or Class 8 environment without contamination risk?
Sterilisation tolerance: Will the material maintain dimensional stability and mechanical properties through the intended sterilisation method (EO, gamma, autoclave)?
Traceability requirements: Can the design accommodate lot traceability markings, serialisation, or UDI compliance without compromising structural integrity?
IEC 60601 alignment: For electromechanical devices, are creepage and clearance distances designed in from the start, or retrofitted under time pressure?
FVG's engineering team engages at the design transfer stage, reviewing component geometry, material specifications, and process parameters before tooling is committed. This upstream involvement routinely prevents the costly design-change cycles that derail programmes during validation.
Medical-grade material selection involves more than choosing a biocompatible polymer. The relevant considerations include:
ISO 10993 biocompatibility requirements for patient-contact applications
Lot-to-lot consistency from approved medical-grade resin suppliers
Processing behaviour under cleanroom injection moulding conditions
Compatibility with downstream assembly and test processes
FVG maintains approved vendor lists for medical-grade materials and manages incoming material inspection as part of its ISO 13485 quality management system.
Injection moulding for medical devices is not the same discipline as industrial injection moulding. The tolerances are tighter, the documentation requirements are more extensive, and the environment in which moulding occurs is controlled to a degree that most general-purpose moulders cannot match.
FVG operates dedicated cleanroom moulding facilities certified to ISO 14644 Class 7 and Class 8 standards. This is not a partition added to an existing moulding floor. It is a purpose-built environment with controlled particulate levels, positive pressure differentials, gowning protocols, and environmental monitoring integrated into the quality management system.
The practical benefits for procurement teams evaluating a moulding partner include:
Reduced contamination risk: Particulate control at Class 7 (ISO 14644) limits particles of 0.5 micron or larger to a maximum of 352,000 per cubic metre. For devices with patient-contact components, this level of environmental control is a regulatory expectation, not a differentiator.
Process validation documentation: FVG's moulding processes are validated through Installation Qualification (IQ), Operational Qualification (OQ), and Performance Qualification (PQ) protocols, producing the documented evidence required for regulatory submissions.
Tooling ownership and maintenance: FVG manages tooling in-house, which means change control, preventive maintenance schedules, and tooling records are part of the same quality system as the moulding process itself.
Medical device components frequently require tolerances in the range of ±0.05mm or tighter for components such as microfluidic channels, catheter hubs, connector housings, and diagnostic cartridge bodies. Achieving these tolerances consistently at volume requires process capability data (Cp/Cpk) established during validation, not assumed from nominal tool dimensions.
FVG's moulding capability covers a range of medical-grade polymers including polycarbonate, ABS, polypropylene, and speciality resins selected for biocompatibility and processing stability.
Modern medical devices are increasingly electromechanical. A diagnostic device that was once a purely mechanical assembly now incorporates a PCB, firmware, sensors, a display, and a wireless communications module. Each of these elements introduces its own regulatory and quality requirements, and the integration of all of them into a finished device is where many programmes encounter their most complex challenges.
FVG's electronics manufacturing capability spans the full electronics value chain within a single facility and quality system.
FVG's SMT assembly lines are configured for the high-mix, low-to-medium volume profile typical of medical device programmes. Key characteristics include:
IPC-A-610 compliance: The industry standard for acceptability of electronic assemblies, with Class 3 workmanship standards applied for medical and high-reliability applications
Component-level traceability: Every component placed on every board is traceable to its source reel, lot number, and date code, providing the audit trail required under ISO 13485 and FDA 21 CFR Part 820
ESD-controlled environments: Electrostatic discharge control protocols throughout the assembly area, protecting sensitive components from latent damage that can cause field failures
Automated optical inspection (AOI): Post-reflow AOI on every board, with X-ray inspection available for BGA and hidden-joint components
The transition from populated PCB to finished medical device involves cable harness assembly, mechanical integration, labelling, and often firmware loading and configuration. Each of these steps requires documented work instructions, operator qualification records, and first-article inspection evidence.
FVG manages the complete box build process, integrating moulded enclosures, electronic sub-assemblies, and mechanical components into finished devices ready for functional test. Because moulding and electronics are performed within the same facility and quality system, the interface between the two is managed internally rather than at a vendor boundary.
When moulding, electronics, and assembly are under one roof, design changes propagate across all three simultaneously. With separate vendors, the same change requires three separate ECO processes, three sets of validation records, and three approval cycles.
Testing in medical device manufacturing serves two distinct purposes that are often conflated. The first is quality assurance: confirming that each unit leaving the production line meets its specification. The second is regulatory evidence: generating the documented data that supports a device's conformity declaration, 510(k) submission, or CE marking under EU MDR 2017/745.
These two purposes require different test strategies, and a manufacturing partner who understands only the first will create problems for the OEM on the second.
FVG develops and executes functional test protocols aligned to the device's design verification requirements. For electromechanical medical devices, this includes:
Electrical safety testing to IEC 60601-1 requirements, covering leakage current, dielectric strength, and protective earth continuity
Functional performance testing against the device's intended use specification
Environmental stress screening where required by the design verification plan
In-circuit testing (ICT) for PCB-level fault detection before integration
The test record is as important as the test result. FVG's quality management system generates Device History Records (DHRs) that capture:
Component traceability to lot and supplier
Process parameter records for moulding and reflow
Inspection results at each stage gate
Test data with pass/fail criteria and operator identification
Non-conformance records and disposition decisions
This documentation structure is designed to support both internal quality management and external regulatory audit. For OEMs targeting Class IIb devices, FVG's alignment with FDA 21 CFR Part 820 and ISO 13485 provides the quality system foundation required for market access in North America and international markets.
FVG supports medical device programmes up to Class IIb under its ISO 13485 quality management framework.
Supply chain management for medical devices is not procurement. It is risk management with a bill of materials. The component shortages of 2020 to 2023 made this viscerally clear to OEM procurement teams who had assumed that their contract manufacturer's supply chain was someone else's problem. It is not. Under ISO 13485 and FDA 21 CFR Part 820, the OEM is responsible for the quality of every component in their device, regardless of who sourced it.
An integrated manufacturing partner changes this dynamic. When FVG manages the supply chain, the approved vendor list, incoming inspection protocols, and supplier qualification records are all part of the same ISO 13485 quality system that governs moulding, assembly, and test. The OEM gets a single quality agreement that covers the entire programme, not a patchwork of separate agreements with separate vendors.
Approved Vendor Lists (AVLs): Supplier qualification based on quality system audits, not just price and lead time
Incoming Quality Control (IQC): 100% dimensional inspection or statistical sampling plans for critical components, with records retained in the DHR
Dual-source strategies: For long-lead or sole-source components, FVG works with OEMs to establish approved alternates before they are needed, not during a shortage
Inventory management: Bonded stock, consignment inventory, and buffer stock programmes to protect against supply disruption without inflating the OEM's balance sheet
Component lifecycle management: Proactive identification of end-of-life components and qualification of replacements within the change control process
FVG's manufacturing footprint across Asia and North America provides geographic optionality that pure single-region manufacturers cannot offer. For OEMs serving global markets, the ability to manufacture in a region aligned with the target market reduces logistics complexity, tariff exposure, and time-zone friction in programme management.
This is particularly relevant for medical device OEMs navigating the evolving trade environment, where supply chain regionalisation has moved from a strategic preference to a procurement requirement for many multinationals.
When procurement teams move from shortlisting to due diligence, the questions that separate genuine integration from marketing copy are specific and process-oriented. The following framework covers the areas that matter most for medical device programmes.
Question | What a Strong Answer Looks Like |
|---|---|
What is the scope of your ISO 13485 certification? | Covers all manufacturing processes performed on-site, not just final assembly |
How do you manage design transfer from OEM to production? | Documented process with defined stage gates, IQ/OQ/PQ protocols, and DHF/DMR management |
What is your CAPA process for non-conformances? | Root cause analysis, corrective action verification, and effectiveness review with defined timelines |
Have you been audited by a Notified Body or FDA? | Yes, with documented outcomes and no unresolved major findings |
Does the partner perform all claimed processes on-site, or are any sub-contracted?
What is their current capacity utilisation, and how would your programme be scheduled relative to existing commitments?
Can they demonstrate process capability data (Cp/Cpk) for critical moulding dimensions?
What is their experience with your specific device classification (Class I, IIa, IIb)?
How many of your critical components have a single approved source?
What is your standard lead time for the top ten components by value in a typical medical device BOM?
How do you manage component end-of-life notifications?
FVG welcomes structured due diligence from OEM procurement teams. The depth of a partner's answers to these questions is itself a quality signal: a manufacturer with mature processes answers them readily, with documented evidence. One without them deflects.
FVG's position in the medical device manufacturing market is defined by a combination of capabilities that are individually common but collectively rare: genuine vertical integration, a mature ISO 13485 quality system, cleanroom manufacturing, electronics assembly, and supply chain management under a single roof and a single quality agreement.
For OEM procurement managers evaluating partners, the relevant proof points are:
ISO 13485 quality management system aligned with FDA 21 CFR Part 820, supporting medical device programmes up to Class IIb
ISO 14644 Class 7 and Class 8 cleanroom facilities in Johor, purpose-built for medical-grade injection moulding and assembly
30+ years of engineering expertise across precision moulding, SMT electronics assembly, and electromechanical integration
Multinational manufacturing footprint across Asia and North America, supporting OEMs with global supply chain requirements
Sustainable operations powered by solar energy, supporting OEM sustainability reporting and ESG commitments
High-mix, low-to-medium volume capability matched to the production profile of most medical device programmes, which rarely resemble the high-volume consumer electronics model
The medical device contract manufacturing market is growing at 10.14% annually because OEMs are making a strategic choice: outsource complexity to partners who are built to manage it, rather than managing it themselves across a fragmented vendor base.
FVG is built for that role. If you are evaluating manufacturing partners for a medical device programme, contact FVG's team to discuss your specific requirements, including device classification, production volumes, regulatory markets, and timeline. The conversation starts with your programme, not a capability brochure.